A co-folding model gives you a complex and a confidence number. SATYA gives you the checked coordinates, every finding at its atoms, and a record you can verify later.
SATYA is an independent structural verification and evidence layer for AI-generated protein-ligand complexes. It does not generate structures and it does not change your file. It checks the coordinates against a structural contract with no experimental reference, gives a whole-structure verdict, localizes each finding to specific residues and ligand atoms with its measurement (a distance between two atoms, or a burial value for a buried polar atom), shows every finding in a 3D workbench where a line appears only for a distance measured between two atoms and is the quantity reported, and exports an evidence package bound by file hashes.
Deterministic. Reference-free. Not built by or tied to any generator. Generated is not verified. The verdict decides. The localization explains. The evidence stays checkable.
A confidence score is the generator's belief about its own output. SATYA is an independent check of the coordinates themselves.
Systems such as OpenFold3, AlphaFold3, Boltz-2 and Chai-1 produce candidate complexes together with confidence or ranking signals. Those signals are internal to the model that produced the structure. SATYA sits after prediction, reads only the structure file, and asks a different question: do these coordinates satisfy a structural contract, and if not, exactly where and by how much? The answer is a verdict for the whole structure plus a list of findings, each pointing at named atoms and carrying its measurement.
DeterministicThe same input bytes produce the same findings. No sampling, no learned scoring, no run-to-run drift.
Reference-freeFindings are violations of a structural contract, not deviations from a crystal structure. No experimental reference is loaded during verification.
IndependentSATYA does not generate structures and has no stake in any generator. It reads the file; it never edits it.
Localized and inspectableEvery finding points at atoms and carries its measurement. Nothing is a black-box score.
Hash-bound evidenceEvery input and output in an evidence package is identified by SHA-256, so the result can be checked later, by you or by anyone you hand it to.
Three kinds of finding, never collapsed into one
REFERENCE_BACKED
deviation from a stated experimental reference
REFERENCE_FREE
violation of a structural contract
COMPARATIVE
changed relation under a declared perturbation
Gate decides. Localization explains. Localization never decides. A reference-free finding is a finding, and SATYA's own verdict travels with it every time it is shown.
The 3D evidence workbench
Not a picture of the result. The result itself, open for interrogation.
The workbench at invariant.pro/structure3d is live and runs in your browser on the published cases. Select a finding and the camera frames the atoms it names; the evidence inspector shows the chain, the residue, the ligand atom and the measurement exactly as the verifier reported it. A measured distance is drawn as a line between the two atoms it was measured between, so the line on screen is the quantity in the record, not an illustration of it; a finding centred on one atom, such as a buried polar atom, is shown at that atom with its burial value and no line. Select any atom in the molecule and the inspector lists the findings that involve it.
Findings, at their atomsEach finding names its chain, residue and atoms and carries the producer-reported measurement: a distance between two atoms, or a burial value for a buried polar atom. The MCL1 case renders its three retained interface measurements at 1.70, 1.73 and 1.92 Å.
Evidence inspectorWhat was checked, what was found, what was not localized, and the admitted source records behind every number, readable inside the page.
Comparative viewFor the KRAS cases, the governed comparison result is shown as its own scientific state. It belongs to the comparison, not to the displayed structure, and it is never rendered as a pass or a fail.
A case summary that decides nothingWhat was checked across six claim classes, what was found, what the case does not include. No overall gate decision is shown here: the workbench explains carried findings and decides nothing.
Figures with honest captionsExport a figure and its record. The caption names the case, the finding in focus, the witness atoms and the counts, generated from the same state the picture was taken from.
Share and replayA share link reproduces the case, the finding in focus and the camera in a fresh browser. A local replay package carries the page, its renderer and its cases for offline review.
The page is served from static files and was verified as served, not as a repository copy: the bytes the public receives were downloaded and exercised in a real browser by the live verification, with every served dependency matched to the tested bytes by digest. The workbench's own code requests only its own files; the host adds its analytics script to the served page.
MCL1 co-fold: coordinate-level evidence the generator's summary did not expose.
In the OpenFold consortium's published OpenFold3 example of MCL1 (four MCL1 chains with three ATP molecules and a small-molecule ligand; seed 42, sample 1), OpenFold3's own summary reported has_clash = 0.0 for the model. SATYA's independent, reference-free review passed the model at its whole-structure gate and localized 15 findings, two of them at the protein-ligand interface, including a reported 1.701 Å distance between HIS55 CE1 and ATP O2G. The generator's summary did not expose this coordinate-level evidence. This is one sample of one generator on one target; it does not establish experimental correctness, a generator deficiency, or how often such contacts occur.
Case statement, fixed wording
Reference-free localization evidence, one generated structure
Structure
OpenFold3 example mcl1, seed 42, sample 1, mcl1_seed_42_sample_1_model.cif, sha256 a6687c4e9d54...
Generator summary
has_clash 0.0, one value for the whole model, generator-reported
SATYA verdict
PASS at the whole-structure gate, co-fold verifier, in its own namespace
Localization
15 findings, 13 protein and 2 interface, every one REFERENCE_FREE
Numbering
model-local residue numbering; the mapping to MCL1 sequence numbering is not verified
The two interface findings, producer-reported distances
Finding
Protein atom
Ligand atom
Distance
Kind
cof_014
HIS55 CE1, chain C
ATP O2G, ligand H
1.701 Å
interface clash
cof_014
HIS55 NE2, chain C
ATP O2G, ligand H
1.729 Å
interface clash
cof_015
ARG94 NH2, chain D
ATP O1G, ligand H
1.918 Å
interface clash
Each of these is drawn in the workbench as a line between the two atoms it was measured between. The published case record binds the structure, the finding payload, the frozen Localization V2 document, the OpenFold3 summary file and the workbench page by digest.
Limits that travel with this case
One sample of one generator on one target.
Every finding is reference-free: a violation of a structural contract, not a deviation from an experimental structure.
SATYA's own PASS verdict is stated whenever the interface findings are.
has_clash is the generator-reported summary. It is not the same measurement as a SATYA finding, and neither corrects the other.
Residue numbers are model-local.
Structure: the OpenFold consortium's published OpenFold3 example (OpenFold/OpenFold3, examples/common_examples/mcl1), Apache-2.0. Credit to the OpenFold consortium.
KRAS / MRTX1133: a blinded prospective benchmark, with its confound disclosed rather than minimized.
In a blinded prospective KRAS/MRTX1133 benchmark, SATYA Layer B resolved a large mutation-associated geometric response at residue 96 across 300 generated structures. Conventional per-candidate plausibility checks, namely PoseBusters, MolProbity and SATYA Layer A, advanced the complete candidate population; their frozen advancement decisions therefore did not distinguish the two conditions. A subsequent post-hoc development analysis showed that the original all-heavy distance descriptor was materially confounded by the tyrosine to serine atom-set change, and that the confound is directional: the frozen descriptor placed Y96S farther from MRTX1133, while a construct-invariant descriptor restricted to N, CA, C, O and CB resolved a large response in the opposite direction, with Y96S closer to the ligand. Residue 96 remained the largest construct-invariant response among the 24 pocket residues, although the pocket response was broad rather than site-exclusive, with 12 of 23 non-perturbed pocket residues also exceeding the same threshold. These post-hoc findings do not alter the frozen prospective Layer B result and do not establish experimentally correct adaptation, spatial specificity, or generalization across targets or generators.
Public claim, fixed wording
Post-hoc development analysis
The frozen result is prospective, blind, reference-free and untouched: 30 seeds per arm, 300 of 300 candidates measured, no seed excluded, no experimental coordinate read. The margin is dimensionless, one pooled within-condition standard deviation; raw_shift is the only Angstrom quantity in the decision path. Everything below was computed after the data were seen, under a directive written and hashed before any number was computed, and it was independently verified 26 of 26.
The direction inversion and its exact decomposition
The two descriptors report opposite-signed responses at the same residue on the same 300 candidates. The perturbed arm is bit identical between them, because SER 96's nearest atom to the ligand already is the backbone oxygen. The whole difference lives in the baseline arm, as tyrosine's extra ring reach.
construct-invariant shift
+0.28847977670787817 Å
real backbone approach, Y96S closer
baseline TYR ring reach advantage
-0.73679032313696702 Å
atom-deletion artifact of the all-heavy descriptor
frozen all-heavy shift
-0.44831054642908885 Å
residual against the frozen value 0.00e+00
The frozen number is a real backbone approach plus an atom-deletion artifact of nearly three times its size, with the artifact winning. The decomposition reproduces at zero residual.
Pocket context, descriptive only
residue 96 absolute g
7.112453833498299
rank 1 of 24 pocket residues
largest control, residue 100
3.7662029017007495
median control
1.0247724983974171
controls above delta
12 of 23
spatial context label BROAD_RESPONSE
The panel carries no nulls and no inferential status and cannot alter the primary classification. It says the pocket response is broad rather than site-exclusive. In Angstroms the site's dominance is cleaner than the standardized ranking: 0.2885 Å is three times the largest control shift and about six times the median control.
The conventional lanes on the same 300 candidates
PoseBusters 300 of 300 advanced. MolProbity 300 of 300 advanced. SATYA Layer A 300 of 300 advanced, PASS 300, FAIL 0, REVIEW 0. These are per-candidate physical plausibility gates; Layer B is a condition-level comparative contrast. They were never asked the same question, and what is frozen is the advancement decision, not the underlying scores.
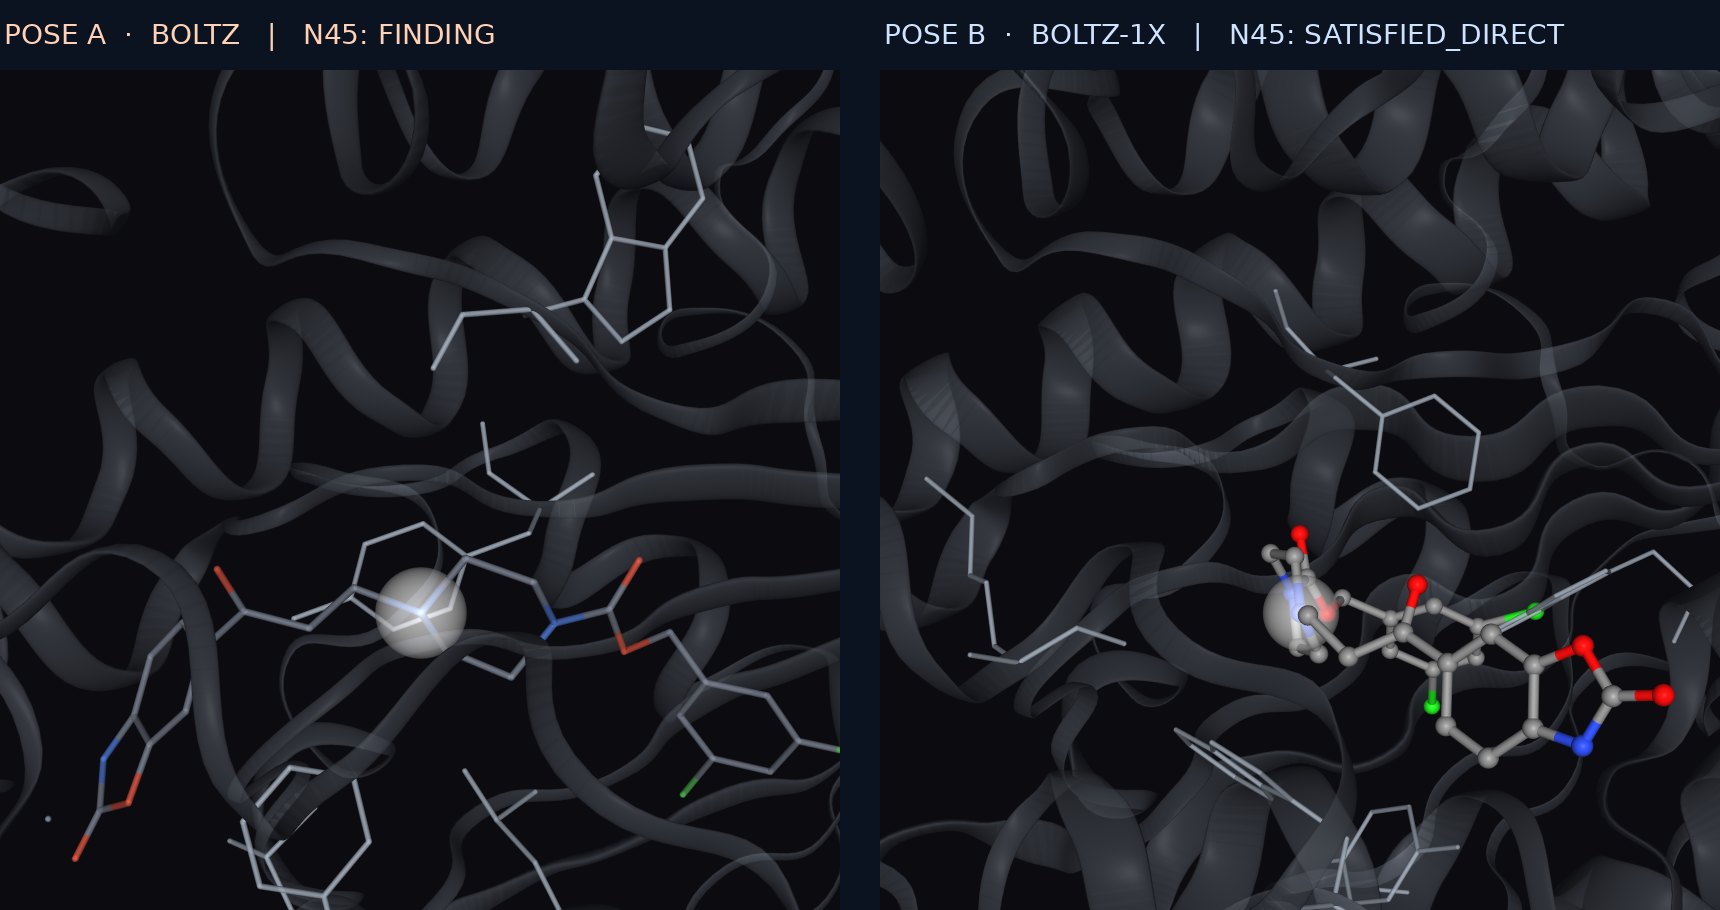
Two generators, one ligand, two different answers. Follow one atom from one pose to the other and read exactly why its evidence changed.
Boltz and Boltz-1x each placed the same ligand in the same protein, and they do not agree. SATYA's W1 channel examines every polar atom of the ligand in each pose and records what it found there: the partner, the burial, the unresolved water. In the workbench you select an atom and switch poses; the same atom stays selected and the inspector shows the recorded reason its outcome changed. Nothing is inferred from a missing flag. Every outcome carries its own evidence.
N45 in the Boltz pose: buried, burial 0.998 against a 0.90 threshold, with no candidate partner within 3.5 Å for any of its roles. That is a W1 finding. N45 in the Boltz-1x pose: a compatible partner for its donor role, the ligand's own O23 at 3.45 Å. No finding. Five of the seven polar atoms that correspond between the two poses change outcome, each with its own recorded reason.
Case statement, values from the published case record
The pair, chosen by rule
System
5s9l, from the V1.2 candidate acquisition
Pose A
Boltz, boltz/637501533__0.cif
Pose B
Boltz-1x, boltz1x/70550355__0.cif
Selection
the first eligible pair under a fixed rule that reads no label, no reference structure and no generator confidence
Polar atoms
7 correspond between the poses; 5 change outcome (O23, N45, N46, O24, O28)
What travels with this case
Every W1 outcome is a structural-contract observation on the submitted coordinates.
The pre-registered confirmation of this diagnostic was NULL (V1_2_PW4B_RUN2): it has not been shown to separate wrong poses from right ones. The complete results are published.
Unflagged means unflagged by this diagnostic, never certified correct.
A SATYA result is not a number in a chat window. It is a case record that binds, by SHA-256, the input structure, the finding payload derived from the verifier's structured output, the source records it was derived from, and the renderer identity it was accepted with. Methods that did not run are named as not run. Findings that could not be localized are listed as not localized. Absence is made visible rather than omitted.
The case recordIdentities, source, generator provenance and input hashes. The structure and the payload are bound by digest before anything is displayed; a mismatch refuses the case rather than rendering it.
The verdict and the findingsA whole-structure verdict in the verifier's own namespace, and a finding list where each entry carries its tier, its atoms, its measurement and its severity scale. Severity scales are per channel and are never compared across channels.
The comparative stateA declared perturbation yields a scientific state, RESOLVED or not, with its effect size, interval and empirical null. It is rendered verbatim. It is never mapped to PASS, SAFE, GOOD or REVIEW.
The frozen protocolDirectives are written and hashed before any number is computed. Prior visibility is disclosed. A result that is not prospective is never described as prospective.
Independent verificationA separate verifier checks what a result artifact certifies, the protected hashes, the gates and the arithmetic re-derived from the recorded inputs, and records an explicit verdict.
Replayable, offlineThe evidence package and the replay package open on a machine with no network connection. What you were shown is what you keep.
How findings are described
A reference-free or comparative finding is never called wrong, an error or incorrect, in the schema or in prose. The generator's own summary values are presented as generator-reported, never as the same measurement as a SATYA finding and never as corrected by one. Model residue numbers are presented as model-local until a mapping is verified. These rules are part of the product, not a disclaimer attached to it.
Pre-registered
The MCL1 open results collection: protocol and sources deposited before a single model was generated.
Invariant is building a public, inspectable record of what SATYA finds on every MCL1 protein-ligand model that can be generated or obtained, published with SATYA's own verdict per model. Every MCL1 protein-ligand structure in the PDB with a small-molecule component, 125 entries, runs through the same accepted route as controls. Published OpenFold3 predictions enter as credited third-party sets. Boltz-2 models are generated under the frozen protocol. Refusals and failures are kept as results; nothing is excluded after the fact.
The frozen protocol, the pinned sources and the index were deposited on Zenodo with a DOI on 2026-10-01, before any generated model was run: zenodo.org/records/23076000, DOI 10.5281/zenodo.23076000. The collection is work in progress. No finding count, rate or comparison is claimed from it until results exist and are recorded.
Third-party structures credited: the OpenFold consortium (OpenFold3, Apache-2.0) for the published MCL1 example; Zonca, Andrea, San Diego Supercomputer Center, DOI 10.5281/zenodo.21926059, CC-BY-4.0, for the 20-prediction OpenFold3 ensemble.
Scope
What this does not claim.
Stated plainly
SATYA does not generate structures, run docking or molecular dynamics, or modify any input file.
SATYA does not claim experimental correctness. A reference-free finding is a violation of a structural contract, and the whole-structure verdict is a structural verdict, not a prediction of binding affinity or biological activity.
The MCL1 result is one sample of one generator on one target. It establishes no rate, no frequency and no statement about any generator.
The KRAS result is a two-arm reference-free comparison. It establishes no experimentally correct adaptation, no spatial specificity and no generalization across targets or generators.
SATYA's threshold checks are sensitive to input coordinate precision at the margins. A three-decimal PDB rendering of a full-precision mmCIF moved results by at most one finding and two violated constraints per model in the conversion control. Format invariance is not claimed.
Verification runs locally on the structure file. There is no upload service on this site, and no structure is sent anywhere by the pages linked here.
Your own structure
The public workbench shows published cases. Your structure is verified on a machine, never on this site.
Nothing is uploaded from the workbench or from this page. Verification of your own co-folded complex runs locally on the structure file, through the same route that produced the published MCL1 case, and comes back as an evidence package you keep and can open offline.
What goes inOne co-folded protein-ligand structure as CIF or mmCIF, with the generator that produced it declared. The file is read; it is never edited.
What runsThe deterministic co-fold verifier on its default thresholds, then localization of every finding to its atoms. No experimental reference is loaded.
What comes backThe whole-structure verdict in the verifier's own namespace, the findings with their measurements, the 3D workbench opened on your case, a figure with its record, and a delivery record binding every file by SHA-256.
What an unsupported input getsA refusal with its reason, not a result: for example a structure with insertion codes or with no protein residues. A refusal is kept apart from a finding and from a machinery failure.
The package reports whatever the verifier finds, a passing verdict included; it is not a promise of a clean result and it does not establish experimental correctness. To arrange a verification of your own structure, write to me.
If your team decides which generated complexes to advance, I would like to understand how you decide today.
Where the decision is made, what evidence it rests on, what it costs when it is made on a structure that should not have advanced, and what an independent, checkable record would have to show you. The public evidence above is the starting point; the workbench is open; the MCL1 collection is pre-registered. Write to me directly.